reached required accuracy - stopping structural energy minimisation
[ctan@baifq-hpc141 primitive_cell-opt]$ tail OUTCAR
                            User time (sec):       88.516
                          System time (sec):        3.010
                         Elapsed time (sec):       94.030

                   Maximum memory used (kb):      238296.
                   Average memory used (kb):          N/A

                          Minor page faults:        36126
                          Major page faults:          669
                 Voluntary context switches:         1271
0) Quit 9) Back ------------->> 2 +---------------------------- Tip ------------------------------+ Input the K-spacing value for SCF Calculation: (Typical Value: 0.03-0.04 is Generally Precise Enough) ------------>> 0.03 Input the K-spacing value for Band Calculation: (Typical Value: 0.03-0.04 for DFT and 0.04-0.06 for hybrid DFT) ------------>> 0.03 +---------------------------------------------------------------+ -->> (02) Reading K-Path From KPATH.in File. +-------------------------- Summary ----------------------------+ K-Mesh for SCF Calculation: 3 3 3 The Number of K-Points along K-Path No.1: 14 The Number of K-Points along K-Path No.2: 6 The Number of K-Points along K-Path No.3: 18 The Number of K-Points along K-Path No.4: 17 The Number of K-Points along K-Path No.5: 7 The Number of K-Points along K-Path No.6: 7 +---------------------------------------------------------------+ -->> (03) Written KPOINTS File.
General timing and accounting informations for this job:
========================================================

                 Total CPU time used (sec):    22600.775
                           User time (sec):    10925.947
                         System time (sec):    11674.828
                        Elapsed time (sec):     7781.069

                  Maximum memory used (kb):     6086716.
                  Average memory used (kb):          N/A

                         Minor page faults:      6831986
                         Major page faults:          282
                Voluntary context switches:     53436870

PROFILE, used timers:     405
=============================
使用Wannier90计算能带
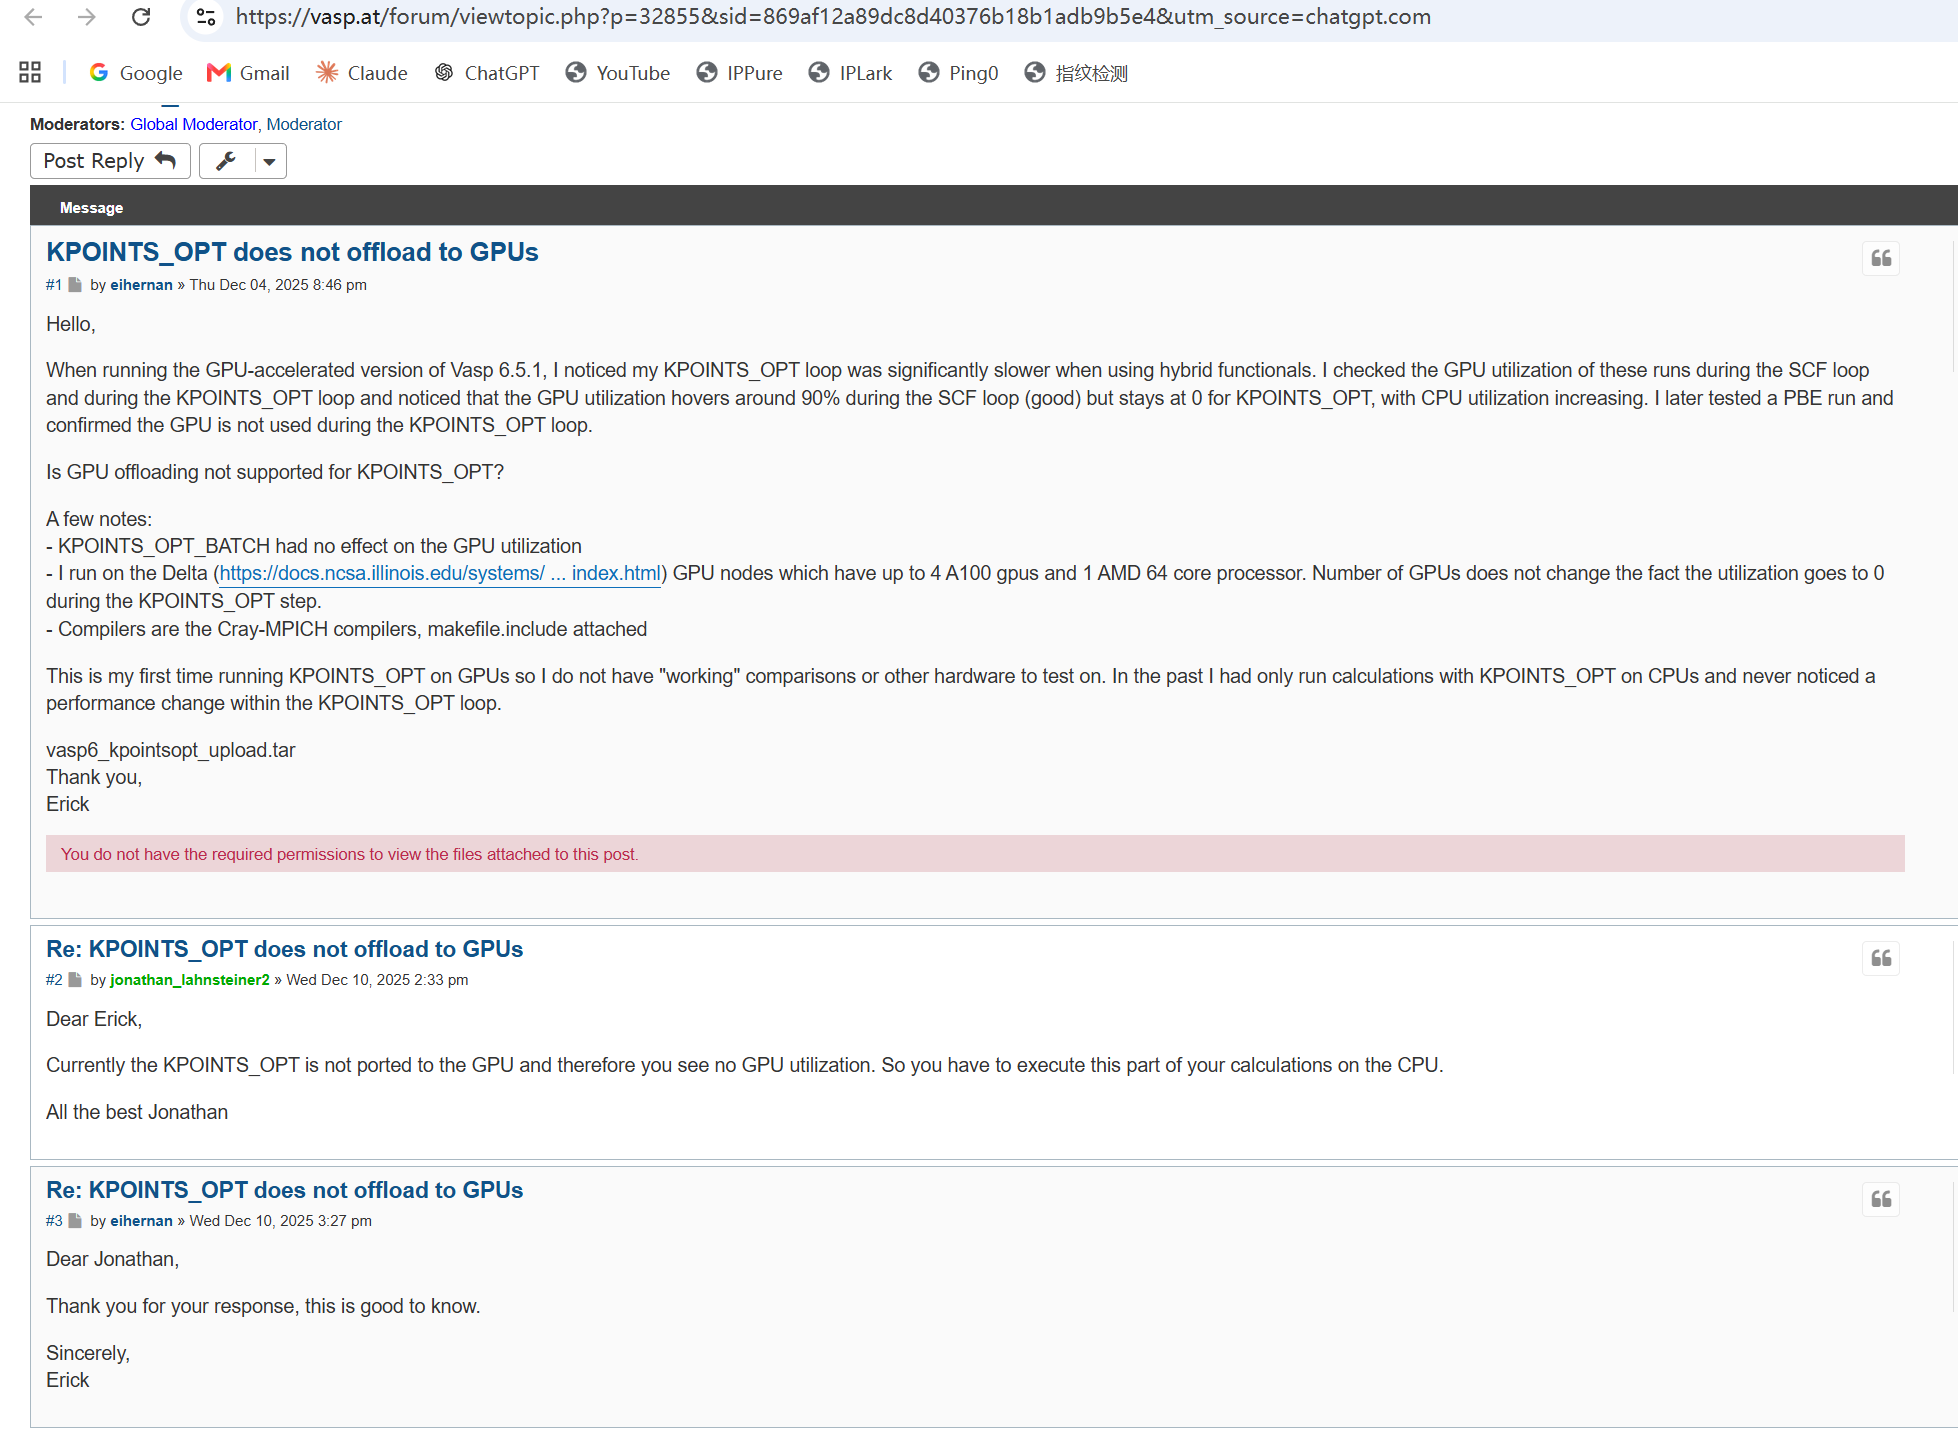
尝试使用wiki推荐的一步方法(VASP 6.6.0 with openACC GPU),KPOINTS中存放SCF所用的均匀k点,然后KPOINTS_OPT中存放高对称点路径。
在HSE06 SCF的部分,GPU满载,但是到了KPOINTS_OPT的部分,GPU空载,CPU 开始工作(开发者给出的答案是“Currently the KPOINTS_OPT is not ported to the GPU.”)整体突然巨慢,这种情况持续了30h+,OUTCAR仍未有任何变化,就不等了。
 ------------>>
21
 ============================ Band Options =======================
 211) Band-Structure
 212) Projected Band-Structure of Only-One-Selected Atom
 213) Projected Band-Structure of Each Element
 214) Projected Band-Structure of Selected Atoms
 215) Projected Band-Structure by Element-Weights
 216) The Sum of Projected Band for Selected Atoms and Orbitals

 0)   Quit
 9)   Back
 ------------>>
211
 -->> (01) Reading Input Parameters From INCAR File.
 +---------------------------------------------------------------+
 |       >>> The Fermi Energy will be set to zero eV <<<         |
 +---------------------------------------------------------------+
 -->> (02) Reading Fermi-Energy from DOSCAR File.
 -->> (03) Reading Energy-Levels From EIGENVAL File.
 -->> (04) Reading K-Path From KPOINTS File.
 -->> (05) Written KLABELS File.
 +---------------------------- Tip ------------------------------+
 |If You Want to Get Fine Band Structrue by Interpolating Method.|
 | You CAN set GET_INTERPOLATED_DATA = .TRUE. in ~/.vaspkit file.|
 +---------------------------------------------------------------+
 -->> (06) Written BAND.dat File.
 -->> (07) Written REFORMATTED_BAND.dat File.
 -->> (08) Written KLINES.dat File.
 -->> (09) Written BAND_GAP File.
-->> (01) Reading Input Parameters From INCAR File. +---------------------------------------------------------------+ | >>> The Fermi Energy will be set to zero eV <<< | +---------------------------------------------------------------+ -->> (02) Reading Fermi-Level From FERMI_ENERGY.in File. -->> (03) Reading Energy-Levels From EIGENVAL File. -->> (04) Reading KPT-Params in the First Line of KPOINTS File. -->> (05) Reading K-Path From KPATH.in File. -->> (06) Written KLABELS File. +---------------------------- Tip ------------------------------+ |If You Want to Get Fine Band Structrue by Interpolating Method.| | You CAN set GET_INTERPOLATED_DATA = .TRUE. in ~/.vaspkit file.| +---------------------------------------------------------------+ -->> (07) Written BAND.dat File. -->> (08) Written REFORMATTED_BAND.dat File. -->> (09) Written KLINES.dat File. -->> (10) Written BAND_GAP File.